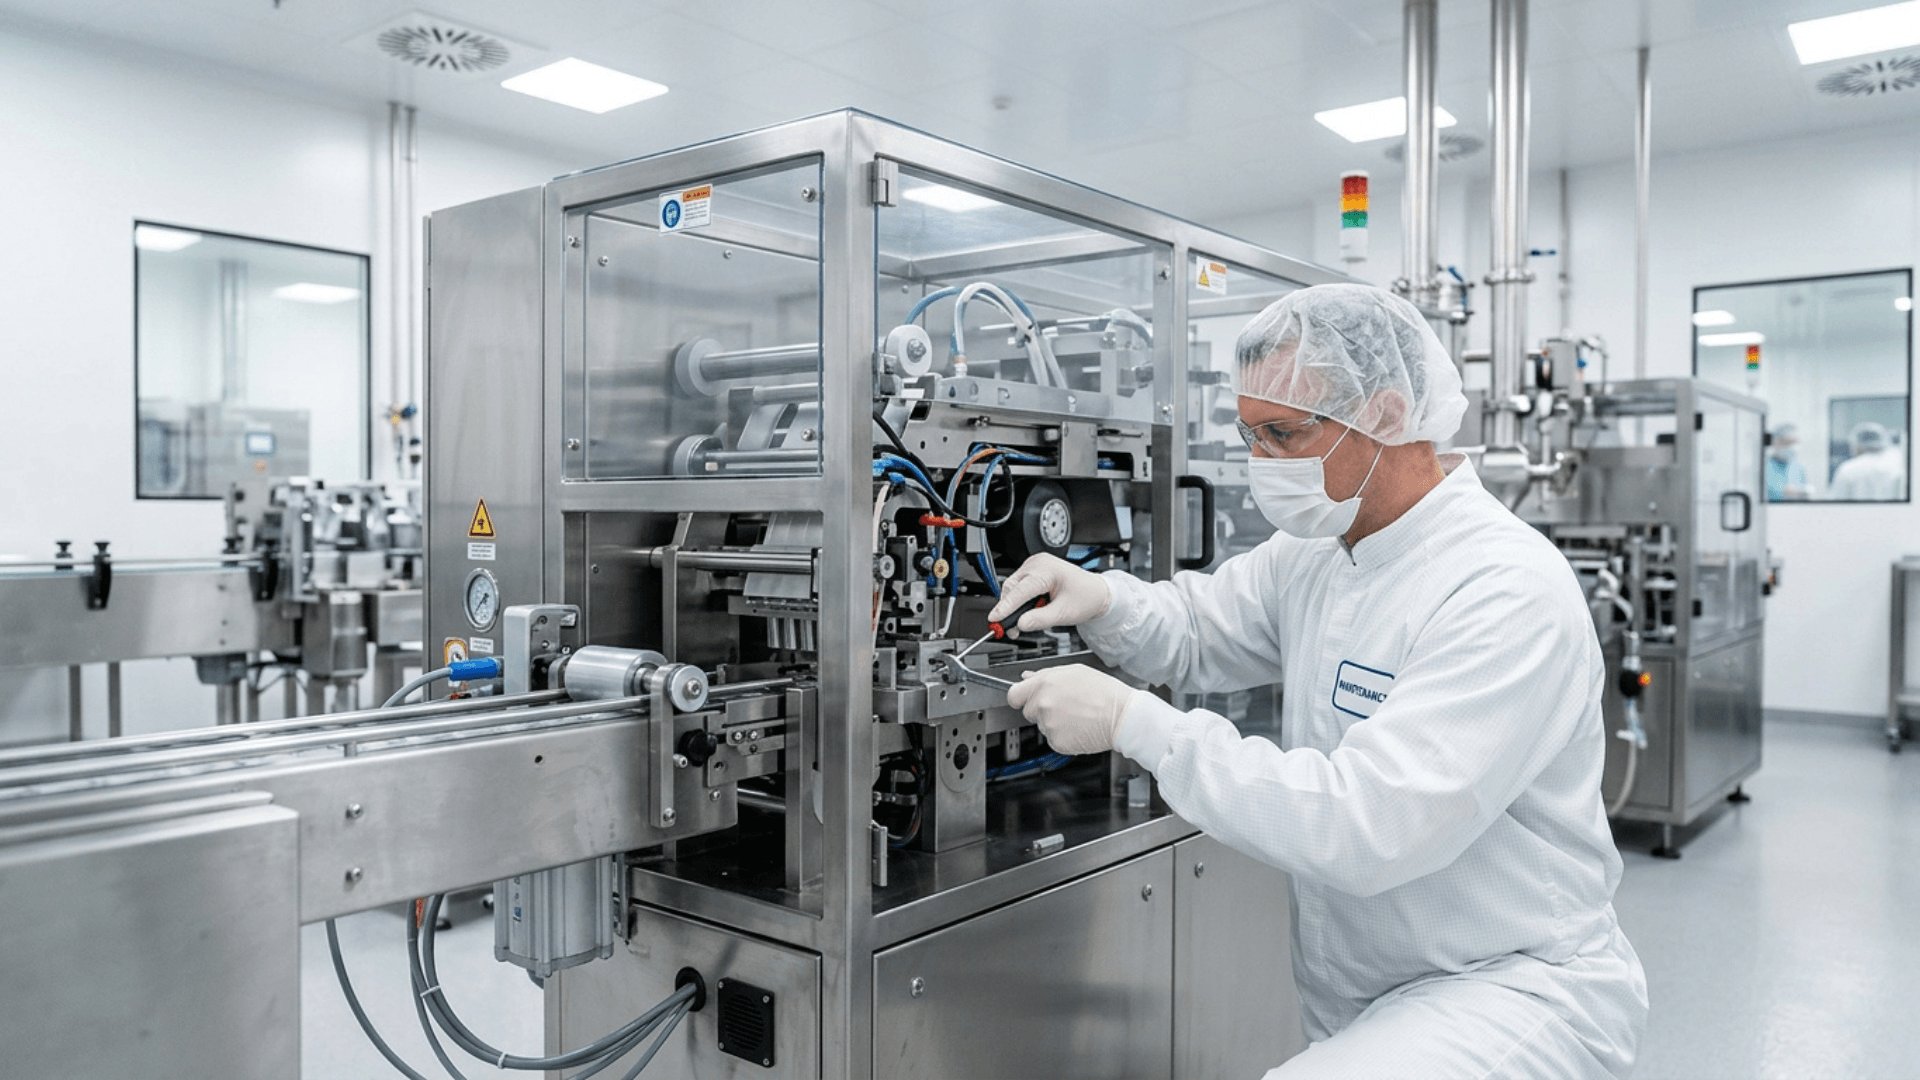
When a technician in a pharmaceutical plant works on a valve or recalibrates a pressure gauge, that intervention becomes part of the history of the batch being produced at that moment. If an AIFA, EMA or FDA inspector later raises a doubt about that batch, the first thing they will ask to see is not the certificate of analysis, but the maintenance log of the equipment involved: who intervened, when, with which spare part, and whether the intervention was closed and verified before production resumed. This is why maintenance in the pharmaceutical industry is subject to the same GMP (Good Manufacturing Practice) standards that govern production, and is treated by regulatory bodies as a Quality function in every respect.
In many plants, however, the maintenance department still works with paper forms, line-level Excel sheets and shared folders updated at different paces by different people. The limitation is not the discipline of the people who work there, but the very nature of these tools, which cannot on their own link an intervention to the batch in production at that moment or flag that a calibration deadline is approaching. When the inspection arrives, reconstructing the relationship between asset, intervention and batch becomes an archival task that can take days, with the concrete risk, in the meantime, of a production stop.
What a GMP inspector is really looking for
A good audit trail does not merely record that an intervention took place. It must make it impossible to modify it retroactively without leaving a trace, preserve the history of every change (who changed what, and when), and make it possible to trace back within minutes from an asset to all the interventions performed in a given period, including any non-conformities found along the way. A CMMS built for regulated environments records this chain at the moment the intervention happens, not after the fact on the initiative of whoever has to prepare for the inspection, and makes it searchable by asset, by batch or by period. Reconstructing the history of a plant thus stops being an archival task and becomes a query.
Validating the software, not just using it
Adopting a maintenance management software in a pharmaceutical company involves a step that goes beyond choosing and configuring the tool: the system must undergo a Computerized System Validation process, following the risk-based approach described in the GAMP 5 guidelines, which classify software according to its criticality and establish which tests — installation, functional, performance — must be run before it can be used in production. Anyone evaluating a CMMS from a Compliance perspective should therefore go beyond simply checking for electronic signatures, and ask the vendor whether it provides supporting validation documentation (requirements matrices, IQ/OQ/PQ protocols, evidence of the tests performed during development) that can reduce the work needed to align with FDA 21 CFR Part 11 and EU GMP Annex 11. A first concrete step in this direction is to move document management out of network folders and into a system that links every document, such as calibration certificates, supplier specifications and operating procedures, directly to the asset and its expiry date. This is how the data integrity required by the ALCOA+ principles (Attributable, Legible, Contemporaneous, Original, Accurate, and then Complete, Consistent, Enduring, Available) becomes a structural property of the system, rather than a commitment to be checked by hand every time.
Why a breakdown is also a contamination risk
Risk classification is the starting point for deciding how stringent a maintenance plan should be: a component in direct contact with the product, such as a valve on an aseptic filling line, requires far more frequent checks than an auxiliary component of the HVAC system serving it. Beyond a certain intervention threshold, which might be the replacement of a critical part on a fermenter, or the overhaul of a gasket on a sterile line, the asset does not simply return to operation: it must be requalified according to the protocol defined for that equipment before production can restart. This is a step that reactive maintenance, managed by trial and error whenever something breaks, struggles to carry out with the required discipline. Maintenance software that supports the implementation of a strategy of preventive maintenance, built on the real life cycles of critical components, makes it possible instead to schedule the intervention and requalification within the downtime window already planned, avoiding both unplanned downtime and the risk of a component failing just while the line is producing.
When a wrong calibration costs an entire batch
What makes calibration critical is not only meeting the deadline, but what happens when an instrument is found out of tolerance at the next check. In that case the correct procedure is not to recalibrate and carry on: an investigation into the deviation must be opened and its retroactive impact assessed on all the batches produced with that instrument since the last valid calibration, which means having to trace quickly every batch involved, not just the date of the last check. Managing calibration schedules on a shared spreadsheet makes this reconstruction slow and error-prone just when speed matters most. Linking every critical instrument to a scheduled maintenance plan, with defined tolerance thresholds, automatic alerts and a complete usage history, reduces the investigation from days of searching to an almost immediate check.
The right spare part, from the right supplier
In the pharmaceutical industry a spare part is not just any interchangeable component. Parts in direct or indirect contact with the product must come from qualified suppliers, with technical documentation and material conformity certificates filed together with the part, because in an audit it is necessary to demonstrate not only that the spare part was installed, but that it came from an approved source. A huge, disorganized warehouse, common in companies that have grown through the successive acquisition of lines and plants, turns this check into an exception that is hard to manage precisely at the moment of a scheduled shutdown, when the critical Clean-In-Place component that is needed cannot be found, or is found but without the documentation that goes with it. Spare parts inventory management that links every component to its qualified supplier, to its minimum reorder thresholds and to the direct and indirect costs of the asset it belongs to turns the warehouse from a risk area into a lever for operational continuity.
The question the system must always be able to answer
What links the audit trail to software validation, and preventive maintenance to calibration and spare parts management, is the same underlying need: being able to answer in real time a very precise question, namely whether, for a given batch, at a given moment, the equipment involved was actually in a state of compliance. An Excel sheet can contain that answer, but only if someone reconstructs it by hand, hoping not to have forgotten anything along the way. A system designed for pharmaceutical Compliance keeps it already linked and verifiable at any time, not only when the inspector arrives.
mainsim, the CMMS and EAM of the Impresoft group, is a horizontal platform used today by companies in manufacturing, food & beverage, healthcare and many other sectors to manage the maintenance of their assets. In the pharmaceutical industry this means linking interventions, deadlines, documentation, calibrations and spare parts in a single verifiable database, integrated with the other business systems already in use.
Those who want to see how other companies have approached this step can start with the success stories collected by mainsim, or request a demo to assess in detail how it applies to their own plant.